Cancer Drug Guidance Shows How FDA Is Cutting Animal Tests Before Human Trials
A May 29 draft lets some oncology biologics rely on one species or a weight-of-evidence assessment instead of three-month primate studies, part of a roadmap launched in 2025.

Key points
- FDA issued draft guidance on May 29 to streamline nonclinical safety studies for certain cancer biologics and conjugated products; comments are due July 30, 2026.
- International ICH S9 guidance allows a cancer drug's chronic effects to be assessed in 3-month studies in one or two animal species; the draft lets some products use rodents only or a weight-of-evidence assessment.
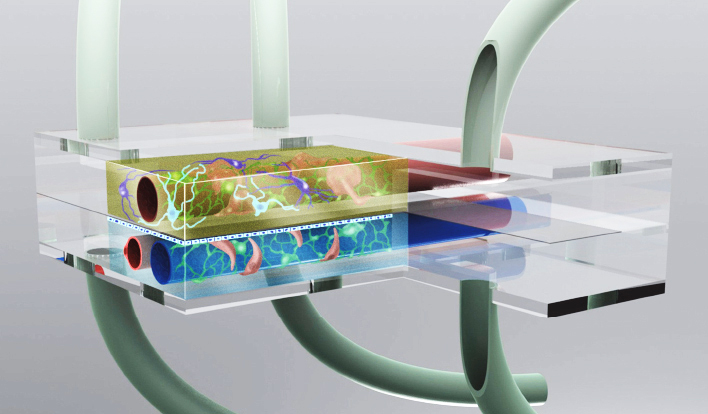
- The draft is part of FDA's April 2025 roadmap to reduce animal testing with new approach methodologies such as organ-on-a-chip systems and computer models.
The Food and Drug Administration on May 29 issued a draft guidance intended to cut what it called unnecessary animal testing in the safety studies done for certain cancer drugs before and during human trials. The document, Oncology Pharmaceuticals: Streamlined Nonclinical Safety Studies for Biologics and Conjugated Products, covers general toxicology studies, with a primary focus on three-month studies. FDA is taking comments until July 30, 2026.
Angelo de Claro, director of FDA's Oncology Center of Excellence, said the recommendations include "eliminating unnecessary animal testing, using a single relevant species instead of two, or replacing animal studies with evidence-based approaches." The draft shows where animal studies sit in drug development, and how FDA is trying to shrink their role.
Why animal studies come first
Drug developers run nonclinical safety studies, many of them in animals, to support clinical development and marketing. For cancer drugs, the reference point is an international guideline from the International Council for Harmonisation known as ICH S9, which FDA implemented in March 2010, and a 2018 question-and-answer document. According to the draft guidance, those documents recommend that assessment of a cancer drug's chronic effects may be based on three-month studies in one or two animal species, which the draft says is sufficient to support continued clinical development and marketing.
Existing guidance already allows some shortcuts. ICH S9 says a repeat-dose study in one rodent species might be enough for some genotoxic drugs, and FDA's 2019 guidance on radiopharmaceuticals generally recommends a risk assessment rather than a general toxicology study, the draft notes.
What the draft would change
The new recommendations depend on the type of product. Animal studies of biologics should use pharmacologically relevant species, meaning animals in which the drug binds its target and has the intended effect; if no such species exists, the safety assessment could rest on a weight-of-evidence risk assessment instead. Biologics active in both rodents and non-rodents could be tested in a single rodent species. For PD-(L)1 blocking monospecific antibodies and CD3 bispecific T-cell engagers, chronic effects may be assessed through a weight-of-evidence review instead of a three-month study. For antibody-drug conjugates whose cytotoxic payload is well characterized and drives toxicity, the three-month study may be done in rodents only.
A weight-of-evidence assessment may draw on the product's own nonclinical and clinical data, literature on the risks of the molecular target, findings for related drugs, and other data such as fit-for-purpose new approach methodologies, or NAMs. CDER's oncology review divisions decide whether that evidence is enough. FDA said the recommendations build on its analysis of general toxicology studies and practices developed during the COVID-19 pandemic to reduce use of non-human primates. Like other guidance, the draft is nonbinding.
The roadmap behind it
The draft is the latest step under FDA's Roadmap to Reducing Animal Testing in Preclinical Safety Studies, published in April 2025. The roadmap describes NAMs as in vitro human-based systems, in silico modeling and other platforms, and cites the figure that over 90% of drugs that appear safe and effective in animals do not go on to receive FDA approval. It notes that the FDA Modernization Act 2.0, passed in late 2022, explicitly authorized non-animal alternatives such as cell-based assays and computer models to support an investigational new drug application.
In an April 20 announcement, FDA said it had met its first-year goals under the roadmap, including draft guidance on reducing or eliminating non-human primate testing for monoclonal antibodies, updated endotoxin-testing guidance and draft guidance expanding weight-of-evidence approaches. The oncology draft supplements, rather than replaces, the ICH S9 documents.
Sources
- FDA Issues Draft Guidance to Cut Unnecessary Animal Testing for Cancer Drugs, U.S. Food and Drug Administration, May 29, 2026
- Oncology Pharmaceuticals: Streamlined Nonclinical Safety Studies for Biologics and Conjugated Products (draft guidance, May 2026), U.S. Food and Drug Administration, May 29, 2026
- Roadmap to Reducing Animal Testing in Preclinical Safety Studies (April 2025), U.S. Food and Drug Administration
- FDA Achieves Year 1 Goals in Reducing Animal Testing in Drug Development, U.S. Food and Drug Administration, April 20, 2026