Blood Irradiator Proposals Show How FDA Classifies Devices Sold Before 1976
Blood irradiators have been cleared through 510(k) for decades without ever being formally classified; FDA's two March 18 proposals show how that gap is closed.

Key points
- FDA proposed on March 18 to classify blood irradiators, which are unclassified preamendments devices, into Class II for preventing transfusion-associated graft-versus-host disease and Class III for preventing metastasis.
- A companion proposed order would require premarket approval applications for the anti-metastasis use by the last day of the 30th month after the Class III classification takes effect.
- FDA said it has cleared 16 blood irradiators under product code MOT, two of them with a metastasis-prevention use.
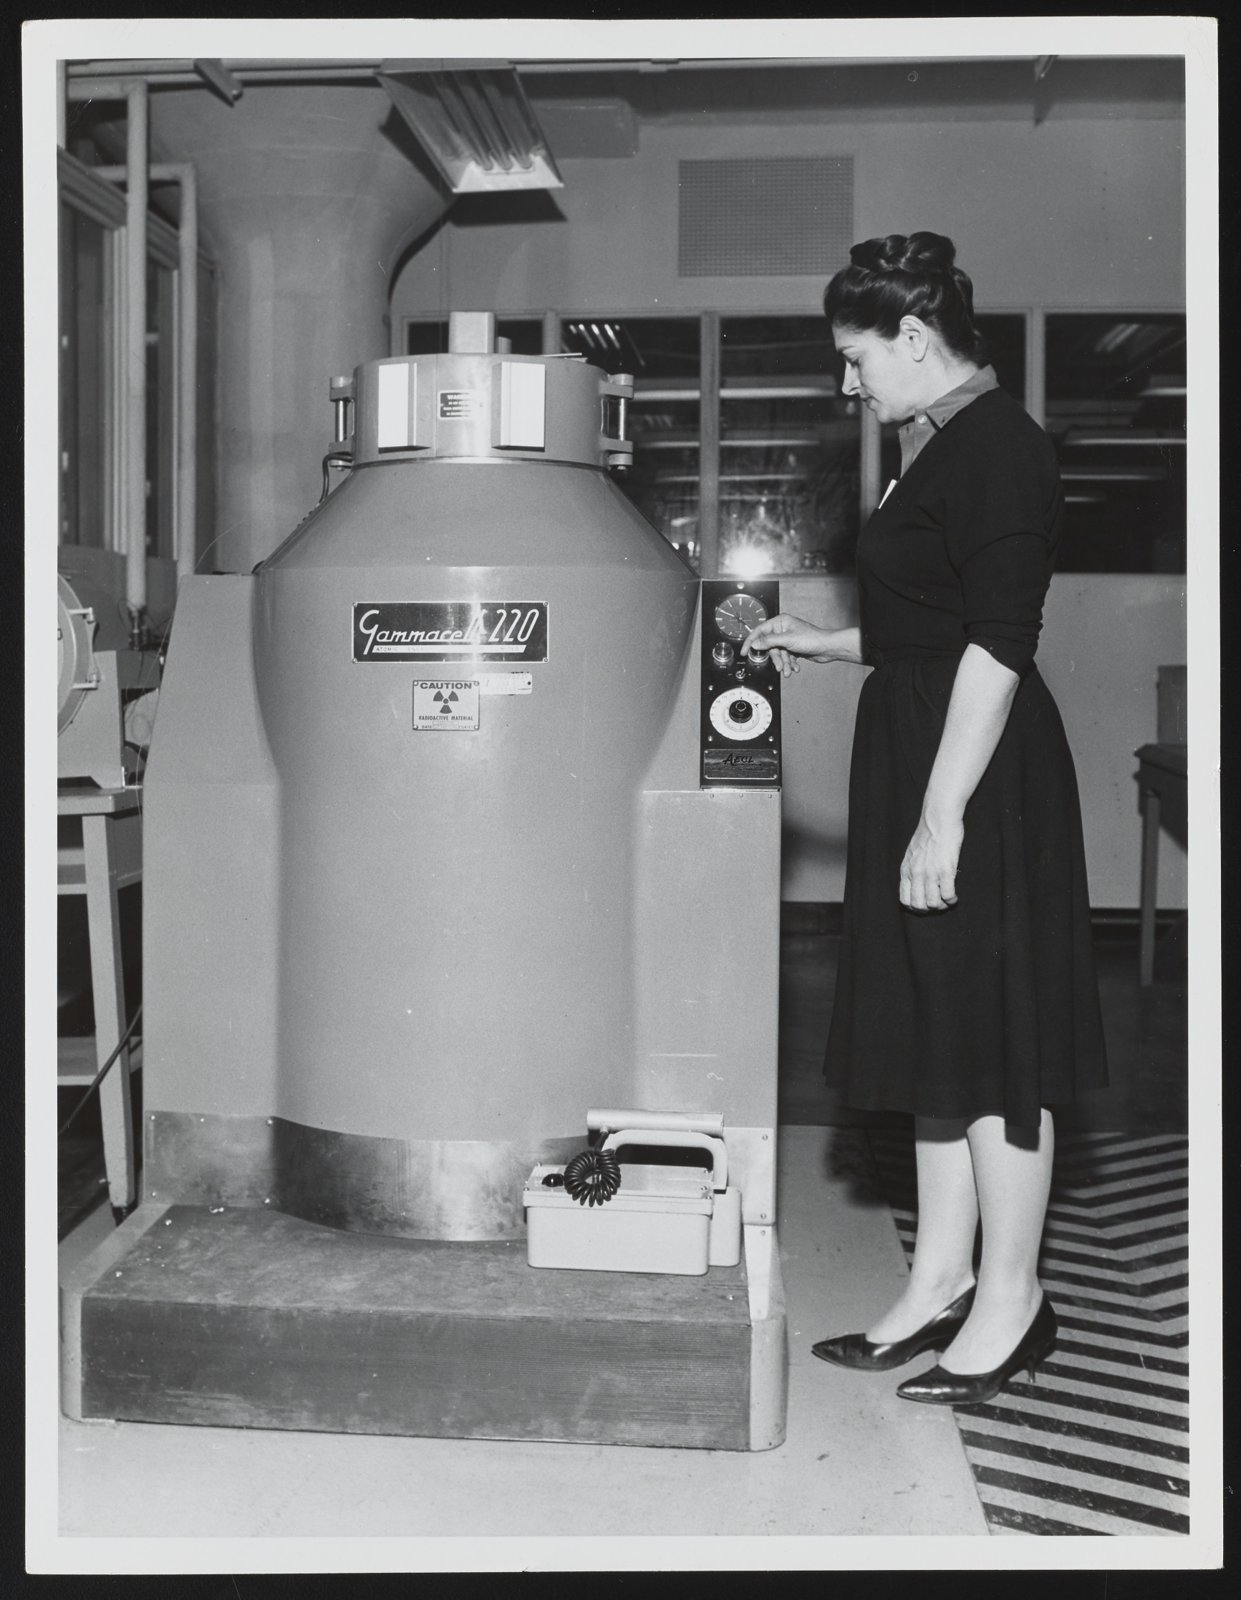
The Food and Drug Administration proposed on March 18 to formally classify blood irradiators, devices that deliver a controlled radiation dose to blood or blood products, which it has regulated for decades without a classification regulation. In a proposed rule, FDA would split the device type by intended use: irradiators that treat blood to prevent transfusion-associated graft-versus-host disease, or TA-GVHD, would go into Class II with special controls, and those used on blood salvaged during cancer surgery to help prevent metastasis would go into Class III.
A companion proposed order would require a premarket approval application for the metastasis use. Comments on both are due May 18. The blood banking group AABB noted that blood irradiators are currently unclassified preamendment devices that FDA regulates through the 510(k) pathway.
Devices that predate the device law
Devices on the market before the Medical Device Amendments of May 28, 1976, are called preamendments devices. Section 513(b) of the Federal Food, Drug, and Cosmetic Act requires FDA to place them in Class I, II or III, and FDA says it has done so for most of them through a set process: a recommendation from a device classification panel, a published proposal and a final regulation.
Blood irradiators were left out. FDA says it cleared the first one for TA-GVHD in 1983 on the basis that it was substantially equivalent to devices sold before 1976, and that in 2005 it cleared the Raycell X-ray Blood Irradiator, the first with a second intended use of preventing metastasis. In all it has cleared 16 devices under product code MOT, two with both uses. Under current practice, marketing a new device in an unclassified type requires a 510(k) clearance.
FDA consulted its Radiological Devices Advisory Panel twice. The 2012 panel recommended Class II for TA-GVHD irradiators. The 2023 panel recommended Class III for the anti-metastasis use, finding insufficient information to show that general and special controls could provide reasonable assurance of safety and effectiveness and that the devices present "a potential unreasonable risk of illness or injury." FDA said it agreed with both panels after reviewing adverse event reports, recall records and published literature. Risks FDA identified for the TA-GVHD devices include improper radiation dose to blood, radiation exposure to the user and electric shock.
How a call for PMAs works
Putting an older device in Class III does not by itself require premarket approval. The proposed order explains that a preamendments device classified into Class III can still be marketed through premarket notification "until FDA issues an administrative order under section 515(b)" requiring premarket approval.
Section 515(b) sets the steps: a proposed order with findings on the risks to be reduced and the public benefit of the device, a comment period, a classification panel meeting, and then a final order or a notice ending the proceeding. If FDA ends it, the law requires the agency to start reclassifying the device under section 513(e), with one exception for banned devices.
Under the proposal, anyone wanting to market a blood irradiator to prevent metastasis would have to file a PMA by the last day of the 30th calendar month after the Class III classification takes effect. The statute lets such devices stay on the market without a PMA until 90 days after a final order or 30 months after the Class III classification takes effect, whichever is later. If no PMA is filed by then and the device is not being studied under an investigational device exemption, it becomes subject to enforcement action, the order says.
Irradiators with an x-ray source are also radiation-emitting electronic products subject to FDA performance standards for cabinet x-ray systems, and those with a sealed radioactive source may fall under Nuclear Regulatory Commission rules, the proposed rule says.
Sources
- Medical Devices; Radiology Devices; Classification of Blood Irradiators (proposed rule), Federal Register (U.S. Food and Drug Administration), March 18, 2026
- Effective Date of Requirement for Premarket Approval Applications for Blood Irradiators Intended To Prevent Metastasis (proposed order), Federal Register (U.S. Food and Drug Administration), March 18, 2026
- REGULATORY UPDATE: FDA Proposes New Classification Rule for Blood Irradiators Used to Prevent TA-GVHD, AABB, March 18, 2026