FDA Warning Letter Cites Zoll Medical Over Quality System and Late Reporting Violations
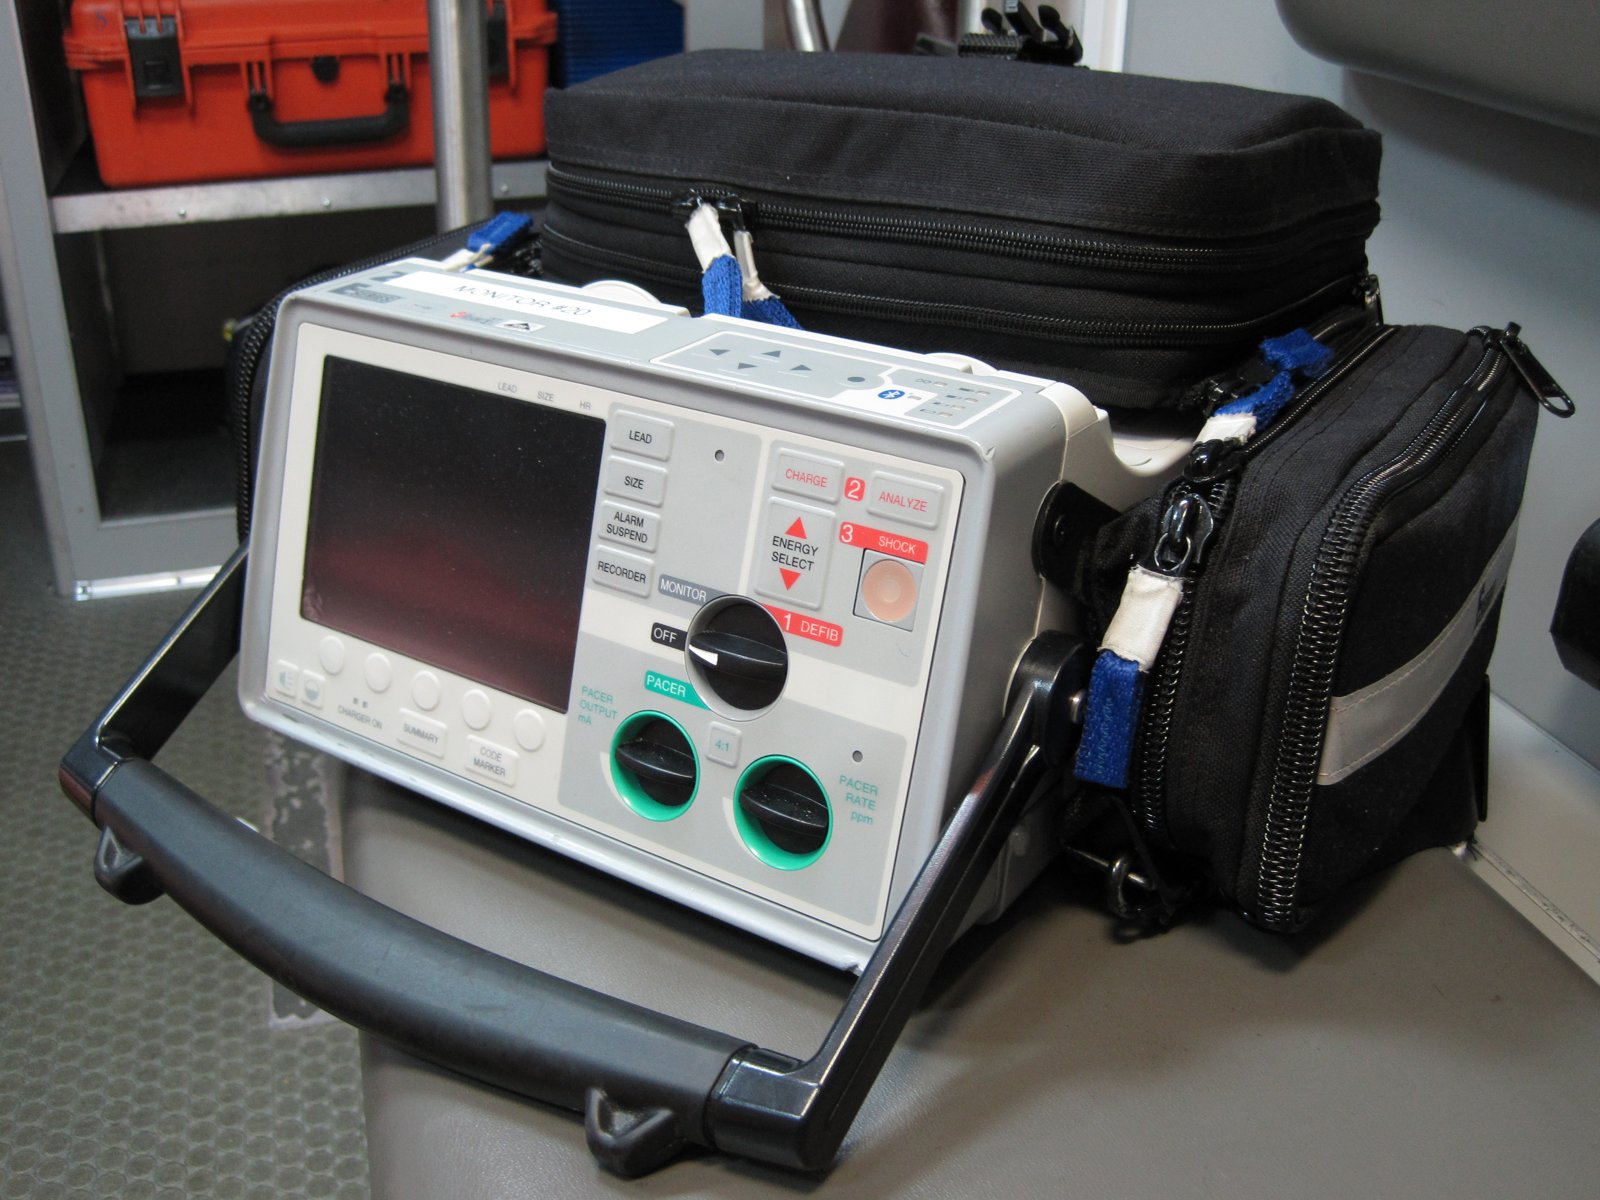
The letter, dated April 30 and now posted by FDA, covers X Series defibrillators, MRI-compatible 731 ventilators, OneStep electrodes and AccuVent sensors made at Zoll's Chelmsford, Massachusetts, site.

Key points
- FDA's device center issued a warning letter to Zoll Medical after a February-April 2025 inspection of its Chelmsford, Massachusetts, facility; the agency's page is current as of June 16.
- The letter cites failures in corrective and preventive action, design validation, risk analysis and supplier controls, plus a medical device report filed months past the 30-day deadline.
- FDA also said Zoll's RescueNet Live software may be a medical device marketed without clearance or approval if it generates its own secondary alarms.
The US Food and Drug Administration has issued a warning letter to Zoll Medical Corporation citing quality system and medical device reporting violations at its Chelmsford, Massachusetts, facility, according to the letter posted on FDA's website. The letter, from the Center for Devices and Radiological Health, is dated April 30, 2026; the agency's page is marked current as of June 16.
The letter follows an inspection conducted from February 27 to April 15, 2025. FDA said the investigator determined that the site manufactures X Series/Advanced automated external defibrillators, Zoll MRI Compatible 731 Ventilators, OneStep electrodes and AccuVent sensors, and that these devices are adulterated because the methods and controls used to make them did not conform to the Quality System regulation. The agency reviewed a series of Zoll responses dated from May 2025 through April 2026 and found several inadequate or impossible to assess.
Corrective action, design and supplier findings
FDA said Zoll did not adequately establish its corrective and preventive action (CAPA) procedure. In one example, a CAPA request opened after a OneStep pediatric electrode was found with burnt tin under the gel was rejected on the grounds that it was a first-time event, even though the company's own record said the nonconformance could cause "arcing and burning a patient." The letter also says complaints about 731 Series ventilators operating near MRI scanners were coded as user errors, no fault found or no product returned, so the CAPA process did not detect the issue; a CAPA was opened on April 9, 2024, and the resulting Class I recall was not initiated until April 30, 2024, the letter says.
The agency cited design validation gaps in the ventilators' MRI compatibility testing, which it said used only a 3T scanner, leaving the labeling without placement guidance for 1.5T scanners. It also said Zoll's risk analysis for X Series defibrillators did not follow the company's own risk management procedure, that complaints about end-tidal CO2 readings and AccuVent ventilation feedback were not escalated to CAPAs, and that two suppliers, including the supplier of RescueNet software, were placed in a category not tied to product safety and effectiveness.
Reporting and software
On medical device reporting, FDA said Zoll became aware on June 20, 2024, of an incident in which a ventilator shut down unexpectedly while operating near an MRI, but the agency did not receive the report until March 10, 2025, beyond the 30-day deadline for malfunctions that could cause or contribute to death or serious injury.
The letter adds that if Zoll's RescueNet Live software analyzes data from a monitoring device and generates a secondary alarm, rather than only displaying alarms, it is a medical device that is adulterated and misbranded because it lacks premarket approval or a 510(k) notification. FDA noted that the inspection was conducted under the Quality System regulation, but that any corrective actions must meet the Quality Management System Regulation in effect since February 2, 2026.
FDA wrote that "Failure to adequately address this matter may result in regulatory action being initiated by the FDA without further notice," including seizure, injunction and civil money penalties. The letter also says that if FDA determines Zoll has violations reasonably related to premarket approval applications for Class III devices, those devices will not be approved until the violations are addressed.
Sources
- Warning Letter: ZOLL Medical Corporation, MARCS-CMS 711320 (letter dated April 30, 2026; page current as of June 16, 2026), US Food and Drug Administration, June 16, 2026