FDA Updates BD Alaris Infusion Set Class I Recall After BD Revises Performance Data
An updated FDA notice adds products after BD said testing showed some sets perform significantly worse than the company had previously disclosed.

Key points
- The FDA updated its Class I recall notice for BD Alaris Pump Infusion Sets on October 5, adding affected products and revised performance measures.
- BD told customers in an August 19, 2026 letter that previously claimed performance for some sets was overestimated, with deviations most common below 1 mL per hour.
- As of August 19, 2026, BD had not reported any serious injuries or deaths associated with the issue, according to the FDA.
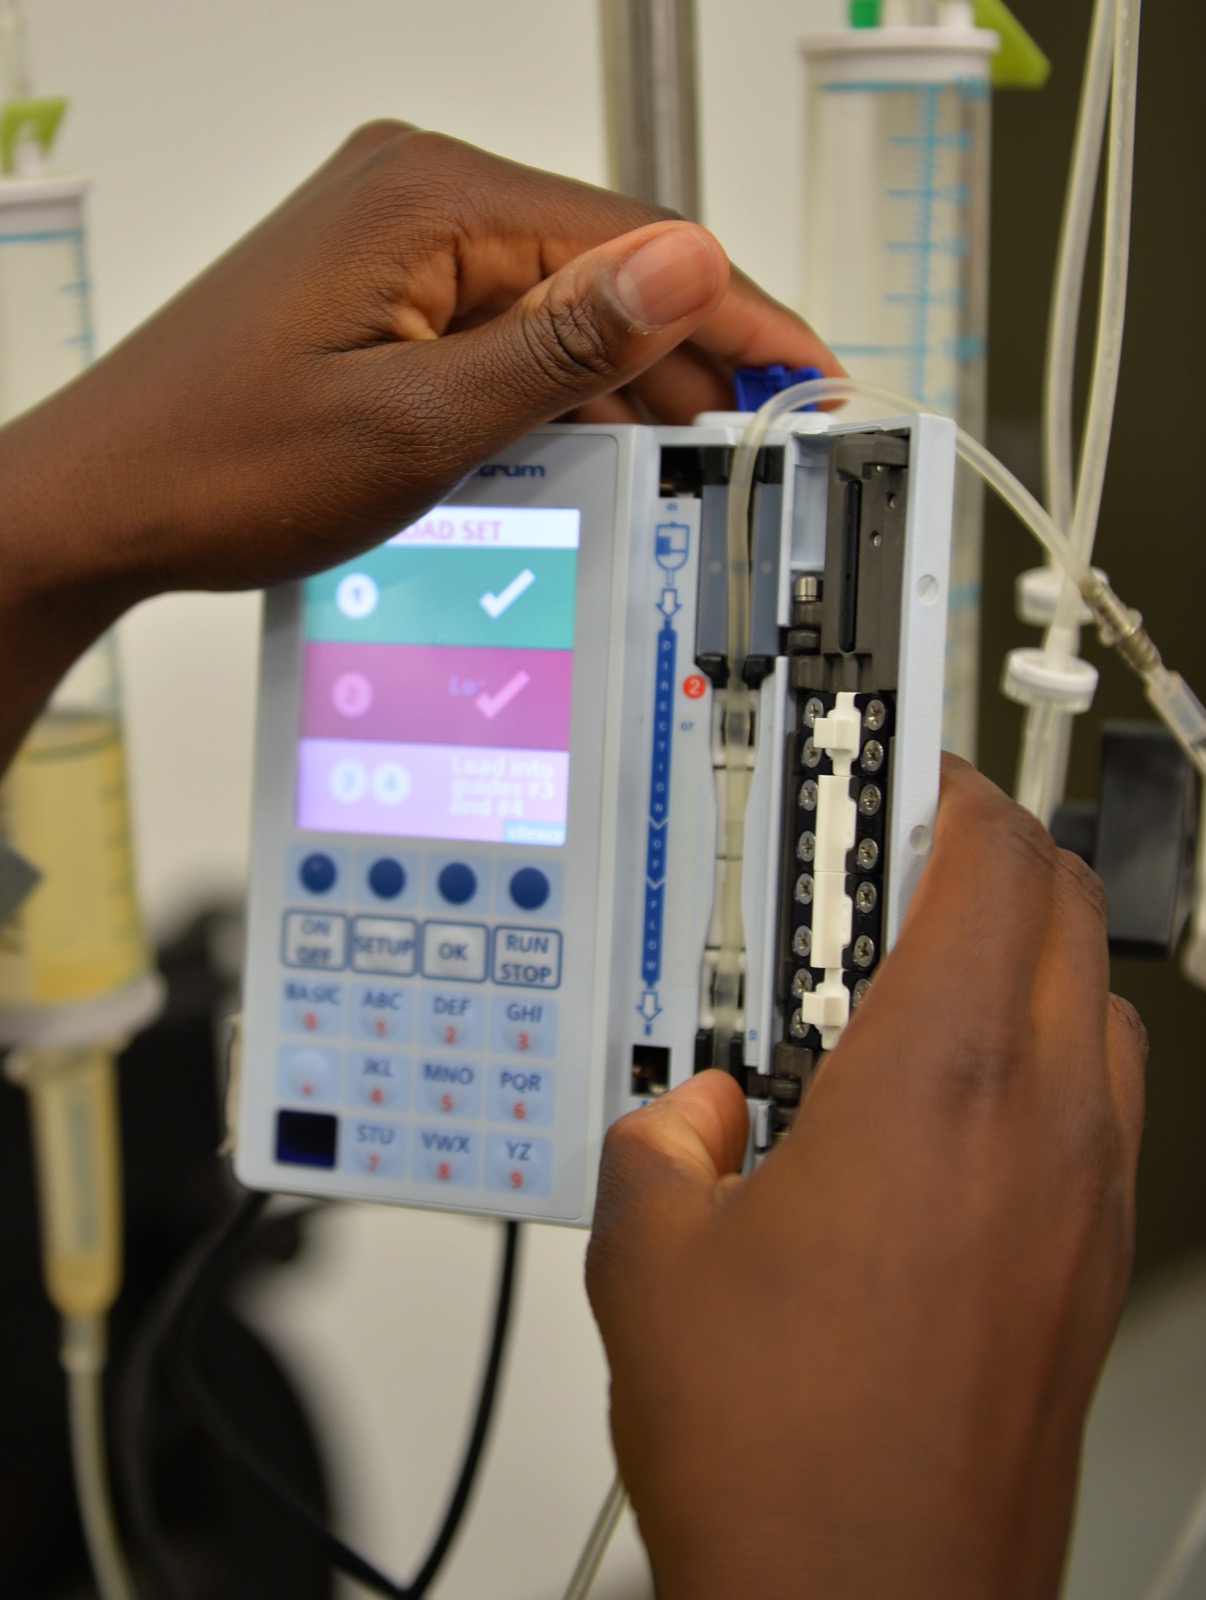
The US Food and Drug Administration on October 5 updated its Class I recall notice for BD Alaris Pump Infusion Sets, adding more affected products and publishing revised performance figures after the manufacturer said some sets perform worse than it had previously disclosed. The FDA notice covers infusion sets used with the BD Alaris Pump Module Model 8100 and is listed as a removal by BD and its subsidiary CareFusion 303, Inc.
The FDA calls a Class I recall "the most serious type." In the notice, the agency states: "This device may cause serious injury or death if you continue to use it."
What BD told customers
According to the FDA notice, BD and CareFusion sent a letter to customers on August 19, 2026. It replaces information from an earlier BD field action, MMS-25-5311, and applies to sets used with the BD Alaris with Guardrails Suite MX software, versions up to v.12.6.
BD said in that letter that "some Alaris Pump sets with restrictive components and designs" perform worse than previously disclosed. Its testing found that "the previously claimed performance was overestimated," and that in some cases "the infusion performance is significantly worse," the FDA notice says.
The deviations occur most often at low programmed rates under 1 mL per hour and at bolus volumes of 5 mL or less, according to the notice. The possible effects it lists are greater under- or over-infusion, delayed occlusion alarms and inaccurate bolus doses. The notice adds: "These examples of performance issues may not be readily detected by users."
For neonates, pediatric patients and critically ill patients, the FDA notice says the issues "could result in serious adverse events or death." As of August 19, 2026, BD had not reported any serious injuries or deaths associated with the issue, according to the agency.
Instructions in the recall letter
The letter, as summarised by the FDA, tells facilities to identify and discard sets listed in an attachment of discontinued Alaris pump sets and to find clinically appropriate alternatives, giving priority to critical-care, neonatal, infant and pediatric areas if supplies are limited. It points users to updated performance disclosures for each set and says that, where performance is not clinically appropriate, an alternative pump such as the BD Alaris Syringe Module or another syringe pump may be used.
If no syringe pumps are available, the letter says the sets "may be utilized with caution, enhanced clinical monitoring," along with the warnings attached to the letter. The FDA notice also reproduces risk-reduction steps from the pump's user manual covering container height, tubing placement over pressure sensors and the handling of occlusions.
A recall that has grown in stages
The October 5 update is the latest step in a recall the FDA has tracked for more than a year. The agency's timeline shows it first issued an early alert on the potentially high-risk issue on July 15, 2025, then updated the communication on September 29, 2025, adding products and performance measures and classifying the issue as a Class I recall. The October 2026 update adds further affected products, current recommended actions and updated performance measures.
The full list of affected devices is published as a separate file linked from the FDA notice. BD has directed customer response forms to its Post Market Quality Operations group and clinical questions to BD Alaris Medical Affairs, according to the notice. Adverse events can be reported to the FDA through its MedWatch program.
Sources
- Infusion Set Recall: BD and CareFusion 303, Inc. Remove BD Alaris Pump Infusion Sets, U.S. Food and Drug Administration, October 5, 2026
- Medical Device Recalls and Early Alerts, U.S. Food and Drug Administration, October 5, 2026