FDA Classifies Conavi Novasight Hybrid Catheter Removal as Class I Recall
The agency upgraded an April 2025 early alert after one reported case in which a coronary imaging catheter's sheath detached and was left in a patient's body before being retrieved.

Key points
- On January 5 the FDA updated its notice on Conavi Medical's removal of Novasight Hybrid catheters to say the action has been classified as a Class I recall, its most serious type.
- According to the FDA, one incident has been reported in which the catheter's sheath detached during use and was left in the patient's body; the full sheath was removed without further injury.
- The FDA recall database lists 685 units in commerce, five lot numbers, and US distribution to Georgia, New York, New Jersey and Ohio, plus Canada and China.
The US Food and Drug Administration has classified Conavi Medical's removal of certain Novasight Hybrid coronary imaging catheters as a Class I recall, the agency's most serious type. The FDA updated its public notice on January 5 to record the classification, nine months after it first flagged the issue in an Early Alert communication on April 18, 2025.
"This device may cause serious injury or death if you continue to use it," the notice says. The action removes devices from where they are used or sold rather than correcting them in place.
One reported sheath detachment
According to the FDA, the recall stems from a single reported incident in which the sheath of a Novasight Hybrid catheter detached during use and was left in the patient's body. "The full sheath was able to be removed from the patient without further injury," the agency said, adding that Conavi has not reported any additional incidents or injuries associated with the issue.
The FDA's recall database entry, recall number Z-1585-2025, gives the manufacturer's reason as "Due to manufacturing issues there is a potential for the catheter sheath to detach." The FDA determined the cause to be process control, according to the entry.
The notice lists the risks the FDA associates with removing a detached sheath from inside a coronary artery, including coronary artery spasm, dissection, perforation, thrombosis, embolism and abrupt vessel closure. The removal process may also bring heart rhythm disturbances, low blood pressure and respiratory insufficiency, and emergency cardiac surgery may be required if retrieval fails, the agency said.
Lots, units and the company's instructions
The affected catheters carry lot numbers 230902, 240202, 240302, 240402 and 240502. The database entry lists the product as the Novasight Hybrid Catheter, model TA-06-0001, cleared under 510(k) number K172258, with 685 units in commerce. Distribution covered Georgia, New York, New Jersey and Ohio in the US, and Canada and China outside it.
Conavi, based in Toronto according to the database entry, sent affected health care providers an urgent product removal notice on March 12, 2025, the FDA said. The firm told customers to stop using the affected catheters, remove them from service and return them to the company, and said the instruction applies to all Novasight Hybrid catheters shipped to the US regardless of expiry date, according to the database entry. Customers were also asked to return an acknowledgement form.
How the device is used
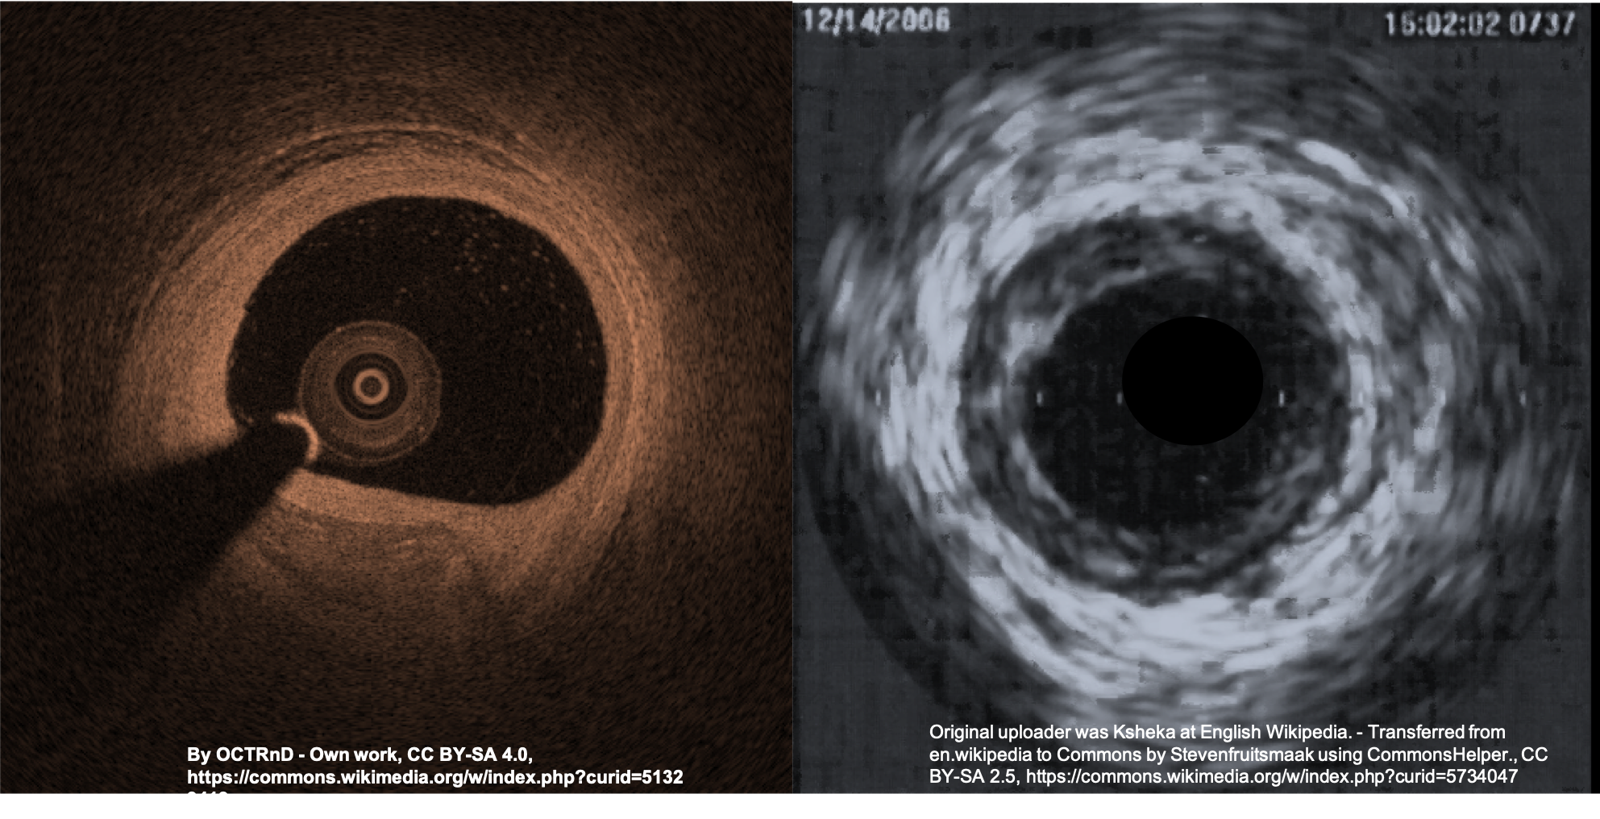
The Novasight Hybrid catheter is part of a system intended for intravascular imaging of coronary arteries, according to the FDA. The database entry describes it as a single-use 3Fr sterile catheter that performs ultrasound and optical coherence tomography imaging at the same time, producing co-registered images of the vessel wall.
The database entry lists the recall as open and classified, meaning the firm had not yet corrected or removed all products. The FDA asks health care professionals and patients to report adverse events or quality problems with the catheters through its MedWatch program.
Sources
- Diagnostic Intravascular Catheter Recall: Conavi Removes Novasight Hybrid Catheters, US Food and Drug Administration, January 5, 2026
- Class 1 Device Recall Novasight Hybrid System (Z-1585-2025), FDA Medical Device Recalls database, April 23, 2025